
翻译:雪球滚滚;出品:Osmosis;编辑:Meducal;
脊髓性肌肉萎缩症
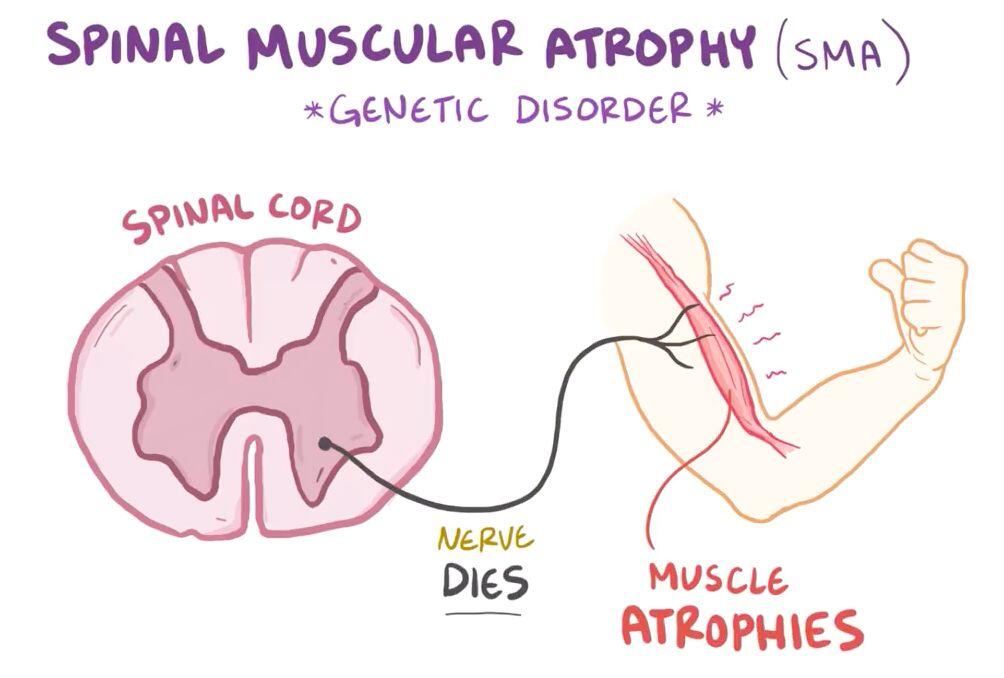
简称SMA,是一种遗传性疾病,会引起脊髓中的神经元细胞过早凋亡,导致其支配的肌肉萎缩、无力。
Spinal muscular atrophy, or SMA, is a genetic disorder** where nerve cells in the spinal cord die prematurely, and this causes the muscles that would normally be controlled by those nerves to atrophy, or wither away, which causes weakness.
α运动神经元
当大脑发出肌肉收缩指令时,上运动神经元将冲动从大脑传到脊髓,接着到达下运动神经元,在这里将来自脊髓的信号传到神经肌肉接头处,即下运动神经元和肌肉细胞相接的地方。
When the brain wants a muscle to contract, it sends a signal through an upper motor neuron, which takes the impulse from the brain to the spinal cord, and then through a lower motor neuron, which goes from the spinal cord to the neuromuscular junction, which is where the lower motor neuron touches the muscle cell.
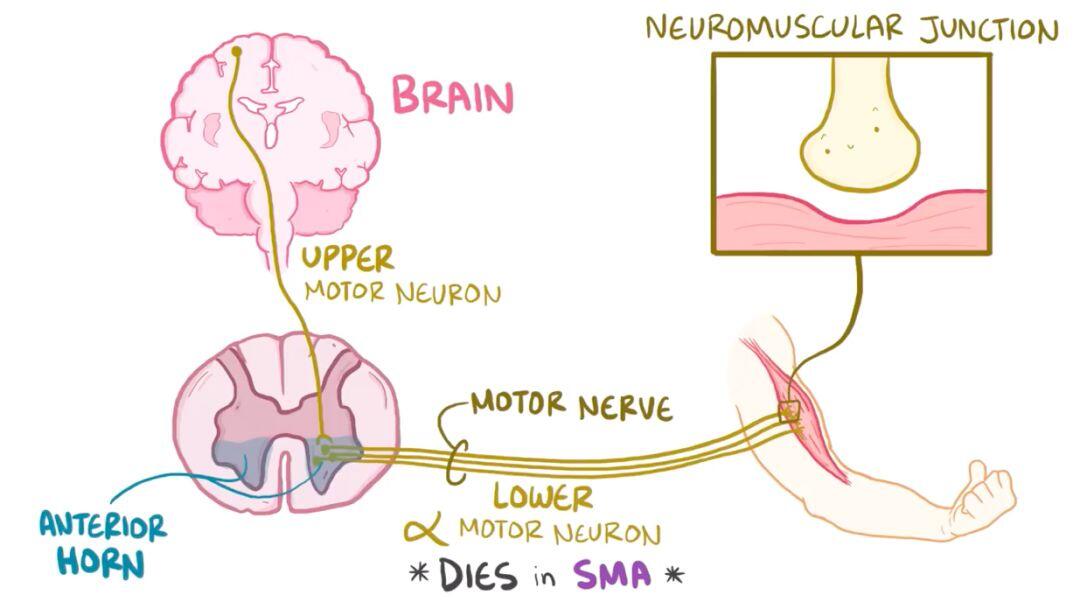
下运动神经元中与骨骼肌的随意收缩有关的叫α运动神经元,这些运动神经元的死亡与SMA有关。其胞体位于脊髓前方的前角,它们的轴突从脊髓投射至其支配的肌肉。这群神经细胞构成运动神经。
The lower motor neurons which cause voluntary contraction of skeletal muscle are called alpha motor neurons, and these alpha motor neurons are the ones that die in SMA. Their cell bodies are located in the anterior horn, or front part, of the spinal cord, and their axons project from the spinal cord all the way to the muscles they innervate. A group of these neurons is called a motor nerve.
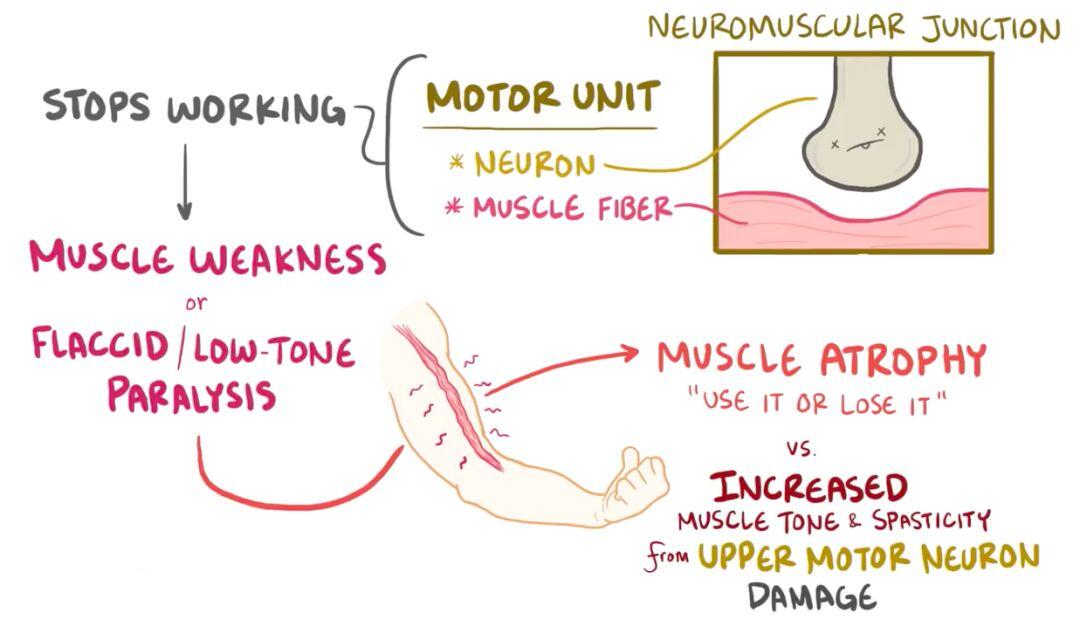
如果下运动神经元死亡或者整条神经受到损害,那么运动单元就会停止工作。运动单元包括运动神经元和其所支配的肌纤维。
If a lower motor neuron dies or if the entire nerve is injured, the motor unit, which includes the neuron and the muscle fibers it innervates, stops working.
根据肌肉纤维无法收缩的程度,可能会出现整个肌肉无力或者说更严重的情况,即弛缓性瘫痪,肌张力降低。
Depending on how many muscle fibers stop contracting, there can be overall muscle weakness or in an extreme situation, a flaccid, or low-tone paralysis.
随着时间的推移,这些失去神经支配的肌肉出现萎缩,很经典的“不用则废”的例子。相形之下,上运动神经元损伤后,会发生肌张力增高、肌肉强直。
This denervated muscle also atrophies over time, a classic example of “use it or lose it”. This contrasts with the increased muscle tone and spasticity that develops after an upper motor neuron is damaged.
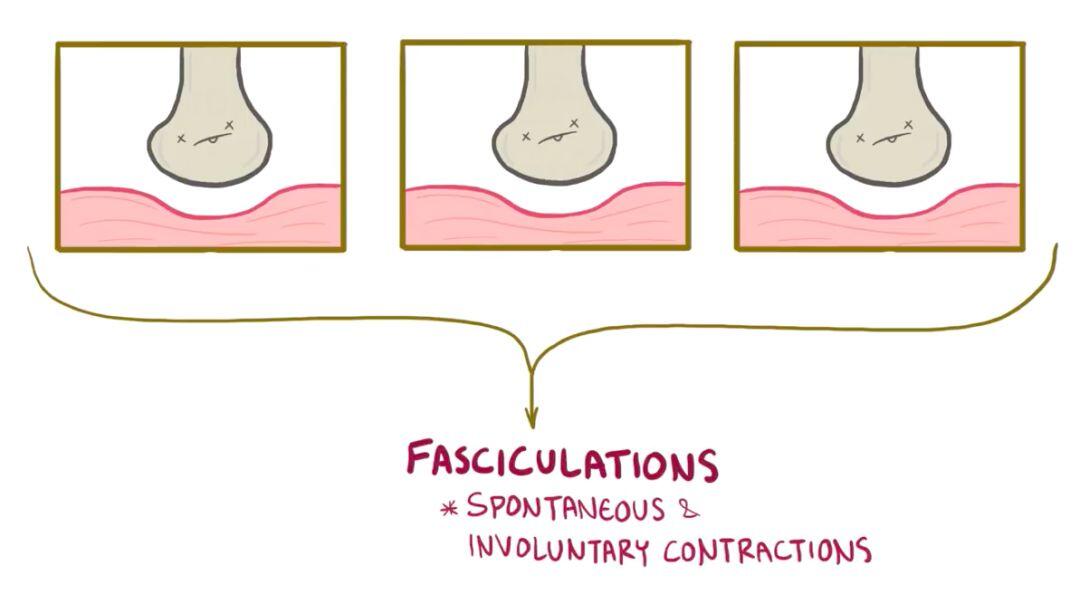
当大量肌纤维受到影响时,就会发生肌束震颤,即自发性肌肉收缩。
When a lot of these muscle fibers are affected, fasciculations can happen which are, spontaneous, involuntary muscle contractions.
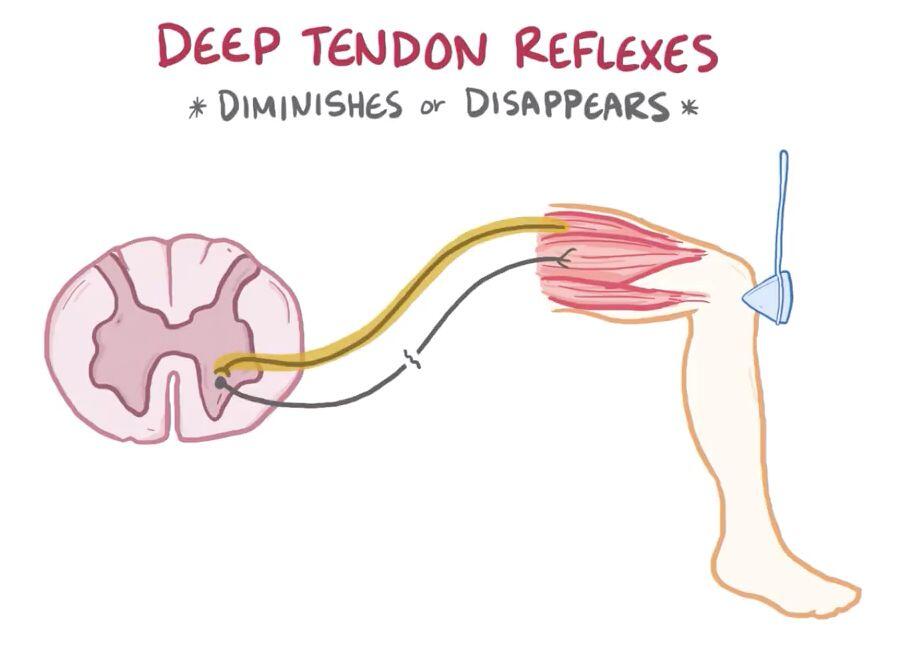
α运动神经元还传导与深反射有关的冲动,例如膝跳反射。如果α运动神经元损伤,深反射会减弱或消失。
Alpha motor neurons also carry the signal for muscle contraction in deep tendon reflexes, like the knee-jerk reflex, and they diminish or disappear when alpha motor neurons are damaged.
SMA类型及症状
来看看SMA的类型及其亚型——
Ia型,又叫先天性SMA,是所有类型中最为严重的一种,甚至在出生前可患病,此时妈妈们会察觉到胎动变弱。
Now, it turns out that there are a few types and subtypes of SMA. Type 1a, congenital SMA, is the most severe of all and it starts even before birth, when mothers may notice decreased fetal movements.
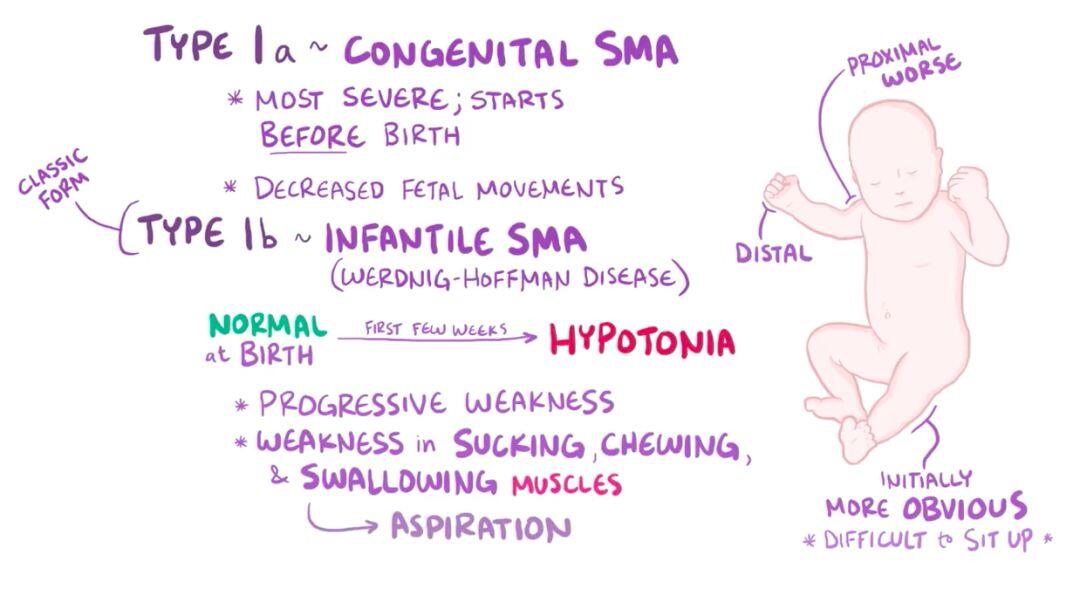
Ib型,也叫婴儿型SMA,或Werdnig-Hoffman病,其典型表现为婴儿在出生后前几周表现正常,随后出现肌张力降低。这些婴儿会出现进展性肌无力,近端肌肉受累比远端严重,最为明显的是小腿,表现为坐起来困难。
SMA type Ib, also called infantile SMA or Werdnig-Hoffman disease, is the classic form where babies often appear normal at birth and then in the first few weeks of life develop hypotonia or low muscle tone. These infants have progressive weakness, which is worse proximally than distally, and is initially more obvious in the legs, making it hard for them to do things like sit up.
同时,患者的吮吸、咀嚼和吞咽功能会受到影响,导致吃奶和进食困难,甚至因无法安全吞咽自身分泌物而吸入异物。胸壁肌肉和膈膜也会受到影响,引起呼吸困难,最终导致呼吸衰竭。因此,这些婴儿大多数仅能存活数年。
They can also have weakness in the muscles involved in sucking, chewing, and swallowing and as a result, they can have difficulty taking milk, eating foods, or even safely swallowing their own secretions which can lead to aspiration. The weakness can also affect the chest wall muscles and diaphragm leading to breathing difficulty and eventually respiratory failure. For these reasons, most of these babies survive only a few years.
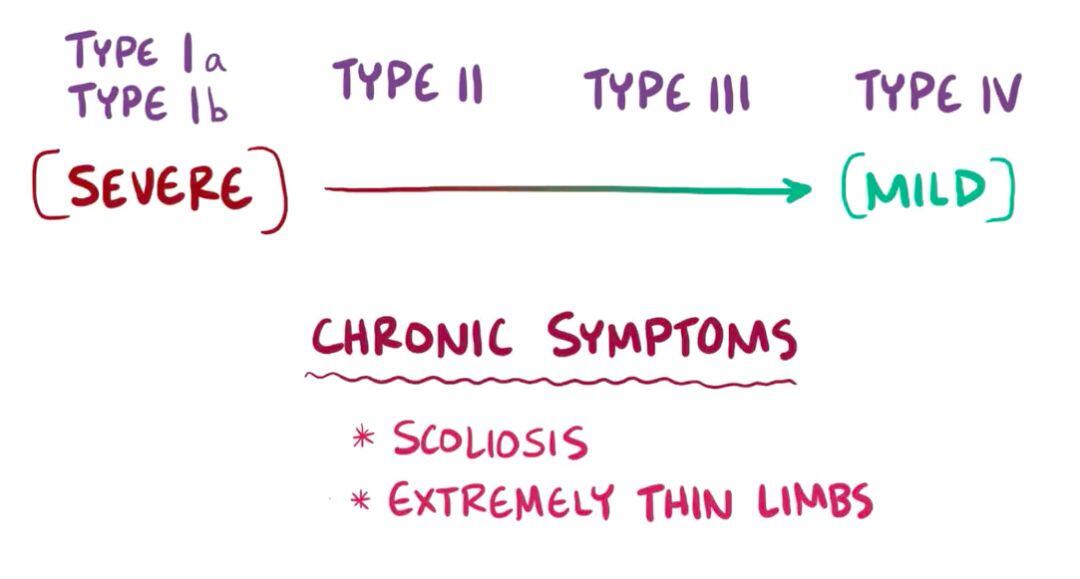
II型、III型和IV型SMA症状更轻,发病更晚。除了肌无力、喂养困难和呼吸困难,长期患有SMA的患者会出现因脊柱肌肉支撑无力导致的脊柱侧弯、及肌肉萎缩导致的四肢瘦弱。
SMA types II, III, and IV are each successively milder and have a later age of onset. In addition to muscle weakness, feeding problems, and breathing difficulties, chronic symptoms of SMA include scoliosis due to poor muscle support of the spine and extremely thin limbs due to muscle wasting.
SMN1基因
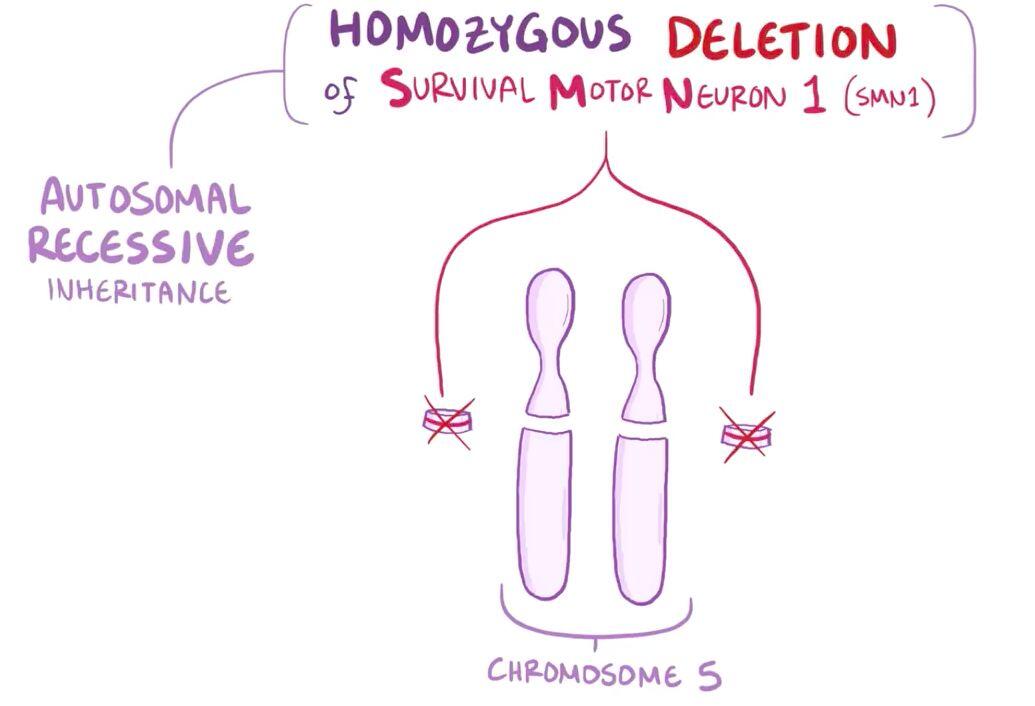
所有不同类型的SMA均因5号染色体纯合性缺失“运动神经元存活”基因(或称SMN1基因)而导致,这是一种常染色体隐性遗传病。
The different types of SMA all result from the same homozygous deletion of the “survival motor neuron” gene or SMN1 gene on chromosome 5, and this is inherited in an autosomal recessive pattern.
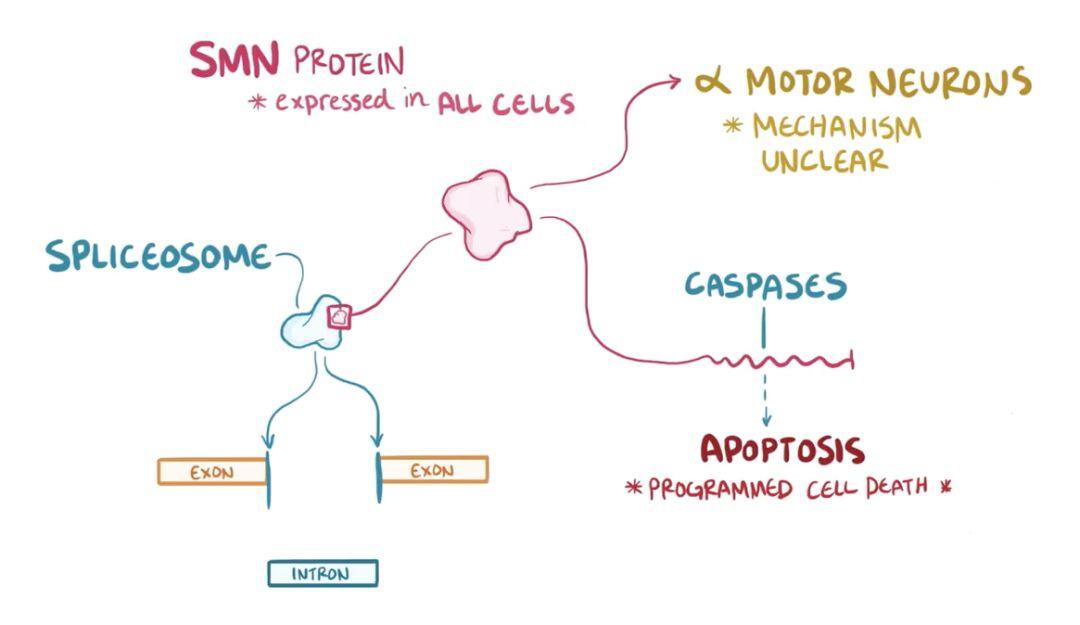
所有细胞都会产生由SMN1基因表达的SMN蛋白,为维持生命所必须的蛋白。从一方面来说,SMN蛋白是剪接体的成分之一,而剪接体就像一台分子机器,作用是切除mRNA前体的内含子。
The SMN protein from the SMN1 gene is expressed in all cells and is required to live. For one, the SMN protein is a component of the spliceosome, a molecular machine that cuts the introns out of pre-messenger RNAs.
SMN蛋白还会阻止胱天蛋白酶的表达,这种酶与细胞凋亡(细胞程序性死亡)有关,因此缺乏SMN,会促进细胞凋亡。SMN蛋白与α运动神经元的关系尤为密切,但是它们之间确切的作用机制还不明确。
SMN also blocks caspases, which are proteins involved in apoptosis, or programmed cell death, so lacking SMN may also enhance apoptosis. SMN protein’s also particularly important for alpha motor neurons, but the exact mechanism relating the protein to the function of those cells is still unclear.
SMN2基因
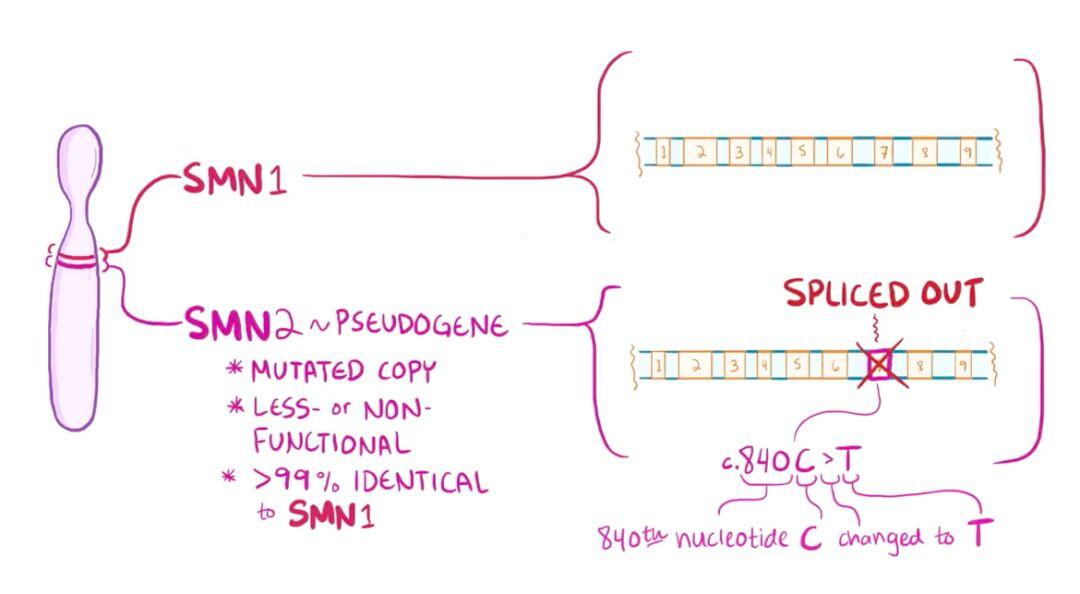
现在,我们可以用遗传学来解释SMA症状严重程度的连续性:这得从SMN2基因说起,这是一种假基因,位于第五号染色体上,旁边就是SMN1。假基因源于进化过程中由拷贝错误产生的突变,与它们对应的基因相比,其功能不全或没有任何功能。
Now, the genetics help explain the continuum of severity that we see across SMA, and it has to do with the SMN2 gene, which is a pseudogene that sits next to SMN1 on chromosome 5. Pseudogenes are mutated copies of genes that arose during evolution, and are less functional or non-functional versions of their counterparts.
假基因:又称“伪基因”。这种基因片段是来自演化过程中的突变累积,有些无法转录,有些则是转录不完全,有些虽然能够转录完成,但也会在转录阶段中途终止作用。
SMN2与SMN1及其相似,但是在外显子7上有明显改变,名叫c.840C>T,即在SMN1中第840个核苷酸是C,而在SMN2中是T。这个微小的突变导致大多数SMN2 mRNA上的外显子7被剪接出来。
SMN2 is more than 99% identical to SMN1, but it has one important change in exon 7, called c.840C>T. And this means that while the 840th nucleotide is a C in SMN1, it’s a T in SMN2. And this tiny mutation results in exon 7 being spliced out of the majority of the SMN2 mRNA.
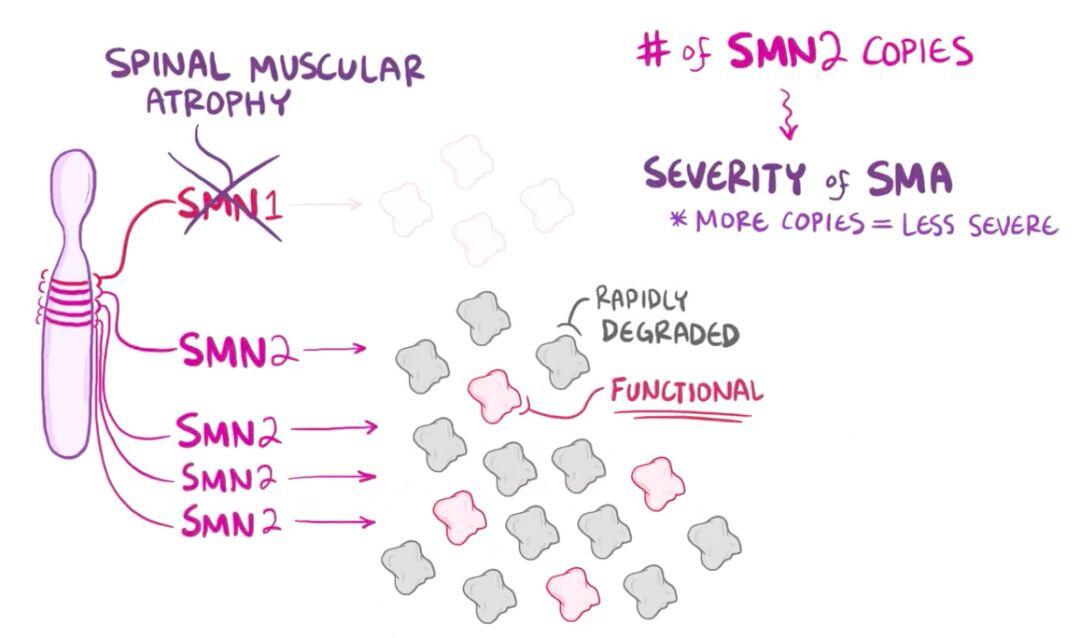
相对于SMN1只产生功能性SMN蛋白,没有外显子7意味着这个SMN2基因产生的SMN蛋白大部分会被迅速降解,只剩下几个全长的功能性SMN蛋白。
Not having exon 7 means this SMN2 gene churns out SMN proteins that mostly get rapidly degraded, with only a couple full-length, functional SMN proteins, relative to SMN1 which churns out only functional SMN proteins.
还有,患者通常都会有多个SMN基因的复制片段,这些可能会导致产生多个SMN2,而这个SMN2拷贝数在人群中实际上变化很大。
Furthermore, often people might have multiple duplications of the SMN genes, which might result in several copies of SMN2 and this SMN2 copy number actually varies quite a bit in a population.
所有这些都很重要,因为SMA患者没有发挥SMN1功能的拷贝片段,而SMN2仍能产生少量的功能性SMN蛋白,所以SMN2拷贝数决定了脊髓性肌萎缩的严重程度:更多的拷贝意味着更多的SMN蛋白和更轻的SMA表型。
Now, all this is important because patients with SMA, have no functioning copies of SMN1, and since SMN2 still makes a small amount of functional SMN protein, the number of copies of SMN2 determines the severity of spinal muscular atrophy. More copies means more SMN protein and a milder SMA phenotype.
举个例子,拥有2个SMN2基因片段的患者会患有婴儿型SMA,而拥有4个SMN2基因片段的患者症状更轻。
So, as an example, a patient with two SMN2 genes might have infantile SMA, whereas a patient with four SMN2 genes might have a milder subtype.
治 疗
SMA的治疗在过去一直是支持疗法,如通过喂食管给婴儿提供营养以及通过给予呼吸支持来缓解肌肉僵硬、增强呼吸肌力量。
Treatment for SMA has historically been supportive, like giving infants nutrition through a feeding tube as well as respiratory support to help with muscle stiffness and strengthen respiratory muscles.
较新的治疗途径是使用Nusinersen。Nusinersen是反义寡核苷酸治疗药物,会与SMN2的mRNA前体结合,阻止其外显子7被移除,从而使SMN2 mRNA得以表达,最终生成较为正常数量的SMN蛋白。
A relatively new therapy for SMA is called Nusinersen. Nusinersen is an antisense oligonucleotide, that binds to the SMN2 pre-mRNA and prevents exon 7 from being removed, which allows the SMN2 mRNA to get expressed, ultimately making a more normal amount of SMN protein.
2016年12月24日,美国食品药品监督管理局(FDA)批准了Biogen和Ionis制药联合开发的脊髓性肌萎缩症(SMA)反义核苷酸治疗药物Nusinersen(商品名Spinraza)上市。
小 结
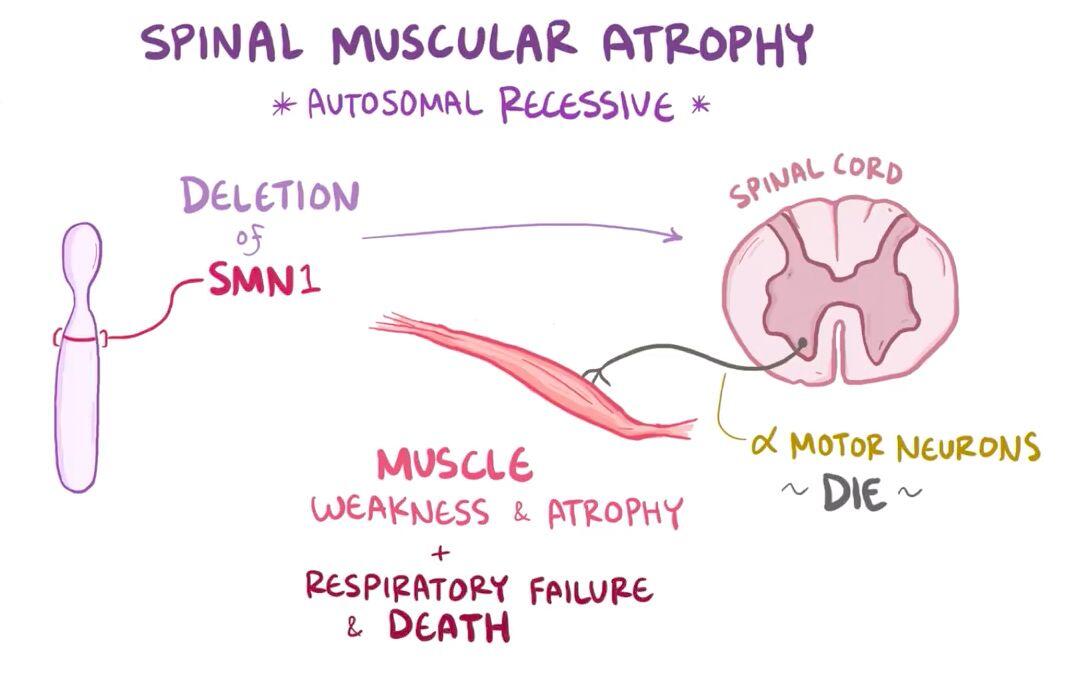
脊髓性肌肉萎缩症,一种常染色体隐性遗传疾病,由SMN1基因缺失导致脊髓中α运动神经元死亡,引起肌无力和肌萎缩;更严重者,会进展为呼吸衰竭和死亡。
All right, as a quick recap, spinal muscular atrophy is an autosomal recessive disorder where a deletion of the SMN1 gene causes alpha-motor neurons in the spinal cord to die, resulting muscle weakness and atrophy and in the more severe cases, respiratory failure and death.
词群索引:https://quizlet.com/_5rmved
👾
MEDUCAL · 医学医生医纪医
(=・ω・=)
